Project 1 - Device Master File
GUIDELINE FOR DRUG MASTER FILES
I. INTRODUCTION
A Drug Master File (DMF) is a submission to the Food and Drug Administration (FDA) that may be used to provide confidential detailed information about facilities, processes, or articles used in the manufacturing, processing, packaging, and storing of one or more human drugs. The submission of a DMF is not required by law or FDA regulation. A DMF is submitted solely at the discretion of the holder. The information contained in the DMF may be used to support an Investigational New Drug Application (IND), a New Drug Application (NDA), an Abbreviated New Drug Application (ANDA), another DMF, an Export Application, or amendments and supplements to any of these.
A DMF is NOT a substitute for an IND, NDA, ANDA, or Export Application. It is not approved or disapproved. Technical contents of a DMF are reviewed only in connection with the review of an IND, NDA, ANDA, or an Export Application.
This guideline does not impose mandatory requirements (21 CFR 10.90(b)). It does, however, offer guidance on acceptable approaches to meeting regulatory requirements. Different approaches may be followed, but the applicant is encouraged to discuss significant variations in advance with FDA reviewers to preclude spending time and effort in preparing a submission that FDA may later determine to be unacceptable.
Drug Master Files are provided for in 21 CFR 314.420. This guideline is intended to provide DMF holders with procedures acceptable to the agency for preparing and submitting a DMF. The guideline discusses types of DMF's, the information needed in each type, the format of submissions to a DMF, the administrative procedures governing review of DMF's, and the obligations of the DMF holder.
DMF's are generally created to allow a party other than the holder of the DMF to reference material without disclosing to that party the contents of the file. When an applicant references its own material, the applicant should reference the information contained in its own IND, NDA, or ANDA directly rather than establishing a new DMF.
II. DEFINITIONS
For the purposes of this guideline, the following definitions apply:
II.1. Agency means the Food and Drug Administration.
II.2 Agent or representative means any person who is appointed by a DMF holder to serve as the contact for the holder.
II.3. Applicant means any person who submits an application or abbreviated application or an amendment or supplement to them to obtain FDA approval of a new drug or an antibiotic drug and any other person who owns an approved application (21 CFR 314.3 (b)).
II.4. Drug product means a finished dosage form, for example, tablet, capsule, or solution, that contains a drug substance, generally, but not necessarily, in association with one or more other ingredients (21 CFR 314.3 (b)).
II.5. Drug substance means an active ingredient that is intended to furnish pharmacological activity or other direct effect in the diagnosis, cure, mitigation, treatment, or prevention of disease or to affect the structure or any function of the human body, but does not include intermediates used in the synthesis of such ingredient (21 CFR 314.3 (b)).
II.6. Export application means an application submitted under section 802 of the Federal Food, Drug, and Cosmetic Act to export a drug that is not approved for marketing in the United States.
II.7. Holder means a person who owns a DMF.
II.8. Letter of authorization means a written statement by the holder or designated agent or representative permitting FDA to refer to information in the DMF in support of another person's submission.
II.9. Person includes individual, partnership, corporation, and association. (Section 201(e) of the Federal Food, Drug, and Cosmetic Act.)
II.10. Sponsor means a person who takes responsibility for and initiates a clinical investigation. The sponsor may be an individual or pharmaceutical company, governmental agency, academic institution, private organization, or other organization (21 CFR 312.3 (b)).
III. TYPES OF DRUG MASTER FILES
There are five types of DMF's:
Type I Manufacturing Site, Facilities, Operating Procedures, and Personnel
Type II Drug Substance, Drug Substance Intermediate, and Material Used in Their Preparation, or Drug Product
Type III Packaging Material
Type IV Excipient, Colorant, Flavor, Essence, or Material Used in Their Preparation
Type V FDA Accepted Reference Information
Each DMF should contain only one type of information and all supporting data. See Section IV.C of the guideline for more detailed descriptions of the kind of information desired in each type. Supporting information and data in a DMF can be cross referenced to any other DMF (see Part V).
IV. SUBMISSIONS TO DRUG MASTER FILES
Each DMF submission should contain a transmittal letter, administrative information about the submission, and the specific information to be included in the DMF as described in this section.
The DMF must be in the English language. Whenever a submission contains information in another language, an accurate certified English translation must also be included.
Each page of each copy of the DMF should be dated and consecutively numbered. An updated table of contents should be included with each submission.
IV. A. Transmittal Letters
The following should be included:
IV. A.1. Original Submissions
a. Identification of submission: Original, the type of DMF as classified in Section III, and its subject.
b. Identification of the applications, if known, that the DMF is intended to support, including the name and address of each sponsor, applicant, or holder, and all relevant document numbers.
c. Signature of the holder or the authorized representative.
d. Typewritten name and title of the signer.
IV. A. 2. Amendments
a. Identification of submission: Amendment, the DMF number, type of DMF, and the subject of the amendment.
b. A description of the purpose of submission, e.g., update, revised formula, or revised process.
c. Signature of the holder or the authorized representative.
d. Typewritten name and title of the signer.
IV. B. Administrative Information
Administrative information should include the following:
IV. B.1. Original Submissions
a. Names and addresses of the following:
(1) DMF holder.
(2) Corporate headquarters.
(3) Manufacturing/processing facility.
(4) Contact for FDA correspondence.
(5) Agent(s), if any.
b. The specific responsibilities of each person listed in any of the categories in Section a.
c. Statement of commitment.
A signed statement by the holder certifying that the DMF is current and that the DMF holder will comply with the statements made in it.
IV. B2. Amendments
a. Name of DMF holder.
b. DMF number.
c. Name and address for correspondence.
d. Affected section and/or page numbers of the DMF.
e. The name and address of each person whose IND, NDA, ANDA, DMF, or Export Application relies on the subject of the amendment for support.
f. The number of each IND, NDA, ANDA, DMF, and Export Application that relies on the subject of the amendment for support, if known.
g. Particular items within the IND, NDA, ANDA, DMF, and Export Application that are affected, if known.
IV. C. Drug Master File Contents
IV. C.1. Types of Drug Master Files
IV. C.1.a. Type I: Manufacturing Site, Facilities, Operating Procedures, and Personnel
A Type I DMF is recommended for a person outside of the United States to assist FDA in conducting on site inspections of their manufacturing facilities. The DMF should describe the manufacturing site, equipment capabilities, and operational layout.
A Type I DMF is normally not needed to describe domestic facilities, except in special cases, such as when a person is not registered and not routinely inspected.
The description of the site should include acreage, actual site address, and a map showing its location with respect to the nearest city. An aerial photograph and a diagram of the site may be helpful.
A diagram of major production and processing areas is helpful for understanding the operational layout. Major equipment should be described in terms of capabilities, application, and location. Make and model would not normally be needed unless the equipment is new or unique.
A diagram of major corporate organizational elements, with key manufacturing, quality control, and quality assurance positions highlighted, at both the manufacturing site and corporate headquarters, is also helpful.
IV. C.1.b.Type II: Drug Substance, Drug Substance Intermediate, and Material Used in Their Preparation, or Drug Product
A Type II DMF should, in general, be limited to a single drug intermediate, drug substance, drug product, or type of material used in their preparation.
IV. C.1.b.(1) Drug Substance Intermediates, Drug Substances, and Material Used in Their Preparation
Summarize all significant steps in the manufacturing and controls of the drug intermediate or substance. Detailed guidance on what should be included in a Type II DMF for drug substances and intermediates may be found in the following guidelines:
Guideline for Submitting Supporting Documentation in Drug Applications for the Manufacture of Drug Substances.
Guideline for the Format and Content of the Chemistry, Manufacturing, and Controls Section of an Application.
IV. C.1.b.(2) Drug Product
Manufacturing procedures and controls for finished dosage forms should ordinarily be submitted in an IND, NDA, ANDA, or Export Application. If this information cannot be submitted in an IND, NDA, ANDA, or Export Application, it should be submitted in a DMF. When a Type II DMF is submitted for a drug product, the applicant/sponsor should follow the guidance provided in the following guidelines:
Guideline for the Format and Content of the Chemistry, Manufacturing, and Controls Section of an Application.
Guideline for Submitting Documentation for the Manufacture of and Controls for Drug Products
Guideline for Submitting Samples and Analytical Data for Methods Validation
IV. C.1.c.Type III: Packaging Material
Each packaging material should be identified by the intended use, components, composition, and controls for its release. The names of the suppliers or fabricators of the components used in preparing the packaging material and the acceptance specifications should also be given. Data supporting the acceptability of the packaging material for its intended use should also be submitted as outlined in the "Guideline for Submitting Documentation for Packaging for Human Drugs and Biologics."
Toxicological data on these materials would be included under this type of DMF, if not otherwise available by cross reference to another document.
IV. C.1.d.Type IV Excipient, Colorant, Flavor, Essence, or Material Used in Their Preparation
Each additive should be identified and characterized by its method of manufacture, release specifications, and testing methods.
Toxicological data on these materials would be included under this type of DMF, if not otherwise available by cross reference to another document.
Usually, the official compendia and FDA regulations for color additives (21 CFR Parts 70 through 82), direct food additives (21 CFR Parts 170 through 173), indirect food additives (21 CFR Parts 174 through 178), and food substances (21 CFR Parts 181 through 186) may be used as sources for release tests, specifications, and safety. Guidelines suggested for a Type II DMF may be helpful for preparing a Type IV DMF. The DMF should include any other supporting information and data that are not available by cross reference to another document.
IV. C.1.e.Type V: FDA Accepted Reference Information
FDA discourages the use of Type V DMF's for miscellaneous information, duplicate information, or information that should be included in one of the other types of DMF's. If any holder wishes to submit information and supporting data in a DMF that is not covered by Types I through IV, a holder must first submit a letter of intent to the Drug Master File Staff (for address, see D.5.a. of this section). FDA will then contact the holder to discuss the proposed submission.
IV. C.2. General Information and Suggestions
IV. C.2.a. Environmental Assessment
Type II, Type III, and Type IV DMF's should contain a commitment by the firm that its facilities will be operated in compliance with applicable environmental laws. If a completed environmental assessment is needed, see 21 CFR Part 25.
IV. C.2.b. Stability
Stability study design, data, interpretation, and other information should be submitted, when applicable, as outlined in the "Guideline for Submitting Documentation for the Stability of Human Drugs and Biologics."
IV. D. Format, Assembly, and Delivery
IV. D.1.
An original and duplicate are to be submitted for all DMF submissions.
Drug Master File holders and their agents/representatives should retain a complete reference copy that is identical to, and maintained in the same chronological order as, their submissions to FDA.
IV. D.2.
The original and duplicate copies must be collated, fully assembled, and individually jacketed.
Each volume of a DMF should, in general, be no more than 2 inches thick. For multivolume submissions, number each volume. For example, for a 3 volume submission, the volumes would be numbered 1 of 3, 2 of 3, and 3 of 3.
IV. D.3.
U.S. standard paper size (8-1/2 by 11 inches) is preferred.
Paper length should not be less than 10 inches nor more than 12 inches. However, it may occasionally be necessary to use individual pages larger than standard paper size to present a floor plan, synthesis diagram, batch formula, or manufacturing instructions. Those pages should be folded and mounted to allow the page to be opened for review without disassembling the jacket and refolded without damage when the volume is shelved.
IV.D.4.
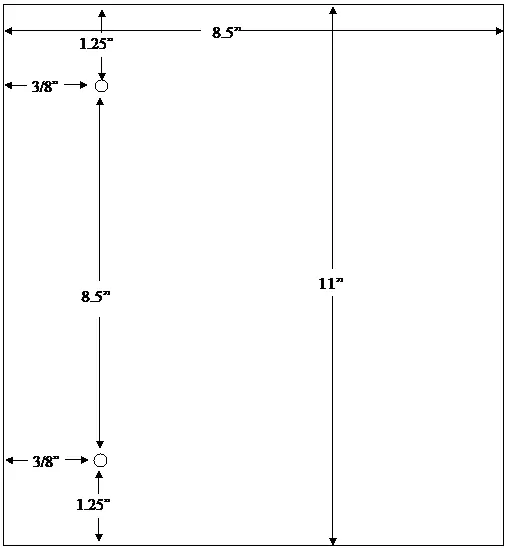
The agency's system for filing DMF's provides for assembly on the left side of the page. The left margin should be at least three fourths of an inch to assure that text is not obscured in the fastened area. The right margin should be at least one half of an inch. The submitter should punch holes 8 1/2 inches apart in each page. See the page measurements shown in the following figure:
IV.D.5. Delivery to FDA
IV.D.5.a.
Drug Master File submissions and correspondence should be addressed as follows:
Drug Master File Staff
Food and Drug Administration
5901-B Ammendale Rd.
Beltsville, MD 20705-1266
IV.D.5.b. Delivery charges to the above address must be prepaid.
V. AUTHORIZATION TO REFER TO A DRUG MASTER FILE
V. A. Letter of Authorization to FDA
Before FDA can review DMF information in support of an application, the DMF holder must submit in duplicate to the DMF a letter of authorization permitting FDA to reference the DMF. If the holder cross references its own DMF, the holder should supply in a letter of authorization the information designated by items 3, 5, 6, 7, and 8 of this section. The holder does not need to send a transmittal letter with its letter of authorization.
The letter of authorization should include the following:
- The date.
- Name of DMF holder.
- DMF number.
- Name of person(s) authorized to incorporate information in the DMF by reference.
- Specific product(s) covered by the DMF.
- Submission date(s) of 5, above.
- Section numbers and/or page numbers to be referenced.
- Statement of commitment that the DMF is current and that the DMF holder will comply with the statements made in it.
- Signature of authorizing official.
- Typed name and title of official authorizing reference to the DMF.
V. B. Copy to Applicant, Sponsor, or Other Holder
The holder should also send a copy of the letter of authorization to the affected applicant, sponsor, or other holder who is authorized to incorporate by reference the specific information contained in the DMF. The applicant, sponsor, or other holder referencing a DMF is required to include a copy of the DMF holder's letter of authorization in the application.
VI. PROCESSING AND REVIEWING POLICIES
VI. A. Policies Related to Processing Drug Master Files
VI. A.1.
Public availability of the information and data in a DMF is determined under 21 CFR Part 20, 21 CFR 314.420(e), and 21 CFR 314.430.
VI. A.2.
An original DMF submission will be examined on receipt to determine whether it meets minimum requirements for format and content. If the submission is administratively acceptable, FDA will acknowledge its receipt and assign it a DMF number.
If the submission is administratively incomplete or inadequate, it will be returned to the submitter with a letter of explanation from the Drug Master File Staff, and it will not be assigned a DMF number.
VI. B. Drug Master File Review
A DMF IS NEVER APPROVED OR DISAPPROVED.
The agency will review information in a DMF only when an IND sponsor, an applicant for an NDA, ANDA, or Export Application, or another DMF holder incorporates material in the DMF by reference. As noted, the incorporation by reference must be accompanied by a copy of the DMF holder's letter of authorization.
If FDA reviewers find deficiencies in the information provided in a DMF, a letter describing the deficiencies is sent to the DMF holder. At the same time, FDA will notify the person who relies on the information in the deficient DMF that additional information is needed in the supporting DMF. The general subject of the deficiency is identified, but details of the deficiency are disclosed only to the DMF holder. When the holder submits the requested information to the DMF in response to the agency's deficiency letter, the holder should also send a copy of the accompanying transmittal letter to the affected persons relying on the DMF and to the FDA reviewing division that identified the deficiencies. The transmittal letter will provide notice that the deficiencies have been addressed.
VII. HOLDER OBLIGATIONS
Any change or addition, including a change in authorization related to specific customers, should be submitted in duplicate and adequately cross referenced to previous submission(s). The reference should include the date(s), volume(s), section(s), and/or page number(s) affected.
VII. A. Notice Required for Changes to a Drug Master File
A holder must notify each affected applicant or sponsor who has referenced its DMF of any pertinent change in the DMF (21 CFR 314. 420(c)). Notice should be provided well before making the change in order to permit the sponsor/applicant to supplement or amend any affected application(s) as needed.
VII. B. Listing of Persons Authorized To Refer to a Drug Master File
VII. B.1.
A DMF is required to contain a complete list of persons authorized to incorporate information in the DMF by reference [21 CFR 314.420(d)]. The holder should update the list in the annual update. The updated list should contain the holder's name, DMF number, and the date of the update. The update should identify by name (or code) the information that each person is authorized to incorporate and give the location of that information by date, volume, and page number.
VII. B.2.
Any person whose authorization has been withdrawn during the previous year should be identified under a suitable caption.
VII. B.3.
If the list is unchanged on the anniversary date, the DMF holder should also submit a statement that the list is current.
VII. C. Annual Update
The holder should provide an annual report on the anniversary date of the original submission. This report should contain the required list as described in B.1., and should also identify all changes and additional information incorporated into the DMF since the previous annual report on the subject matter of the DMF. If the subject matter of the DMF is unchanged, the DMF holder should provide a statement that the subject matter of the DMF is current.
Failure to update or to assure FDA annually that previously submitted material and lists in the DMF remain current can cause delays in FDA review of a pending IND, NDA, ANDA, Export Application, or any amendment or supplement to such application; and FDA can initiate procedures for closure of the DMF (see Section IX).
VII. D. Appointment of an Agent
When an agent is appointed, the holder should submit a signed letter of appointment to the DMF giving the agent's name, address, and scope of responsibility (administrative and/or scientific). Domestic DMF holders do not need to appoint an agent or representative, although foreign DMF holders are encouraged to engage a U.S. agent.
VII. E. Transfer of Ownership
To transfer ownership of a DMF to another party, the holder should so notify FDA and authorized persons in writing. The letter should include the following:
- Name of transferee
- Address of transferee
- Name of responsible official of transferee
- Effective date of transfer
- Signature of the transferring official
- Typewritten name and title of the transferring official.
The new holder should submit a letter of acceptance of the transfer and an update of the information contained in the DMF, where appropriate. Any change relating to the new ownership (e.g., plant location and methods) should be included.
VIII. MAJOR REORGANIZATIONOF A DRUG MASTER FILE
A holder who plans a major reorganization of a DMF is encouraged to submit a detailed plan of the proposed changes and request its review by the Drug Master File Staff. The staff should be given sufficient time to comment and provide suggestions before a major reorganization is undertaken.
IX. CLOSURE OF A DRUG MASTER FILE
A holder who wishes to close a DMF should submit a request to the Drug Master File Staff stating the reason for the closure. See Section IV.D.5.a for the address.
The request should include a statement that the holder's obligations as detailed in Section VII have been fulfilled.
The Agency may close a DMF that does not contain an annual update of persons authorized to incorporate information in the DMF by reference and a list of changes made since the previous annual report. The holder will be notified of FDA's intent to close the DMF.
IMPORT LICENSES – DEVICE MASTER FILE (DMF)
The registration process in India for imported devices is a largely a re-assessment of their existing approval, utilizing the same documentation and evidence required in the reference country. This being said, the CDSCO will pay very close attention to the predicate device comparison section of the application. It will be important for manufacturers to carefully select the predicates compared to and provide sufficient evidence for equivalence.
Device Master File
- It is the detailed form of information regarding the product which is submitted to the relevant Governmental authority and basically which is intended for the incorporation into the Regulatory Dossier
- It is a Device reference source submitted to the FDA as defined by the 21 CFR 814.
Manufacturing license
- With respect to a country, any regulatory authorization required to manufacture one or more products or classes of product as granted by the relevant governmental authority
Indian Regulatory Authority Name
- Central Drugs Standard Control Organization (CDSCO) , Under Ministry of Health & Family Welfare
The Global Market size of medical devices-
Indian Medical Device Market-
With a CAGR of 15.8%, the Indian medical device market increased from USD 2.02 billion (INR 13,130 crores) in 2009 to USD 3.9 billion (INR 25,259 crores) in 2015.
In 2015, this represented roughly 1.7% of the market for medical devices worldwide
- The Indian healthcare market, estimated at USD 96.7 billion (INR 6.29 lakh crores) in 2015, receives 4% of its revenue from the sale of medical devices.
- According to industry estimates, the Indian medical device market would expand by 16�GR to reach USD 8.16 bn (INR 53,053 crore) in 2020.
- Now India has become one of the top 20 global medical device markets and the 4th largest medical device market in Asia
Medical device classification based on Risk-
- Class A is for low risk
- Class B for low moderate risk
- Class C for moderate to high risk
- Class D for high risk
Regulatory process
Form No. to apply and grant Manufacturing Licence and Loan Licence for Class A & B Medical devices and IVD devices
Manufacturing license
|
Classification |
Application |
Granted license |
Licensing authority |
|
Class A &B |
Form MD-03 |
Form MD-05 |
State License Authority (SLA) |
Loan license
|
Classification |
Application |
Granted license |
Licensing authority |
|
Class A &B |
Form MD-04 |
Form MD-06 |
State License Authority (SLA) |
Form No. to apply and grant Manufacturing Licence and Loan Licence for Class C & D Medical devices and IVD devices
Manufacturing license
|
Classification |
Application |
Granted license |
Licensing authority |
|
Class C & D |
Form MD-07 |
Form MD-09 |
Central License Authority (CLA) |
Loan license
|
Classification |
Application |
Granted license |
Licensing authority |
|
Class C &D |
Form MD-08 |
Form MD-10 |
Central License Authority (CLA) |
Form No. to apply and grant Manufacturing Licence and Loan Licence for all Class new Medical devices and IVD devices
Application & Approval Form required for New medical device
Manufacturing license(New medical device)
|
Classification |
Application |
Granted license |
Licensing authority |
|
Class A,B, C&D |
Form MD-26 |
Form MD-27 |
Central License Authority (SLA) |
Manufacturing license(New in vitro medical device)
|
Classification |
Application |
Granted license |
Licensing authority |
|
Class A,B,C &D |
Form MD-28 |
Form MD-29 |
Central License Authority (CLA) |
Loan license for new medical devices and In Vitro devices
|
Classification |
Application |
Granted license |
Licensing authority |
|
Class A & B |
Form MD-04 |
Form MD-06 |
State License Authority (SLA) |
|
Class C &D |
Form MD-08 |
Form MD-10 |
Central License Authority (CLA) |
Power of Attorney
- Authorised Agent who has been appointed by the overseas manufacturer for import,sales and distribution of Medical device or IVD devices
- Official representative for Regulatory Compliance
The documents needed to apply manufacturing license for Class A Medical Devices are-
|
Sl no. |
Medical Device |
IVD device |
| 1. |
Cover letter |
|
| 2. |
Constitution details of the firm(of the domestic manufacturer or authorized agent) |
|
| 3. |
The Establishment/Site ownership/Tenancy Agreement |
Not Applicable |
| 4. |
ISO 13285 Certificate or similar QMS Certificate |
|
| 5. |
Site or Plant master file |
|
| 6. |
Device description,intended use,specifications and accessories |
|
| 7. |
Working principle and use of a novel technology |
|
| 8. |
Materials of construction |
Analytical technology,relevant analytes and test procedure |
| 9. |
Labeling and Pack size |
|
| 10. |
Summary of reported adverse events |
Analytical Performance Report |
| 11. |
Essential principal checklist of the medical device |
Essential Principle Checklist of the In Vitro Diagnostic Medical Device |
| 12 |
Undertaking Letter stating that Compliance with QMS |
|
| 13 |
Test License or existing License as per Drugs and Cosmetics Act,1940-For Indian Manufacture only |
|
| 14. |
Fee Challan |
|
| 15 |
Legal Form |
|
The list of documents to be needed to apply manufacturing licence for Class B Medical Device and IVD Medical Device-
|
Sl.no. |
Medical Device |
IVD Device |
|
1. |
Cover Letter |
|
|
2. |
Constitution details of the firm(of the domestic manufacturer or authorized agent) |
|
|
3. |
The Establishment/Site ownership/Tenancy Agreement |
Not Applicable |
|
4. |
ISO 13485 Certificate or similar QMS Certificate |
|
|
5. |
Site or Plant master file |
|
|
6. |
Device Master File |
|
|
7. |
Essential Principle Checklist of the Medical Device |
Essential Principle Checklist of the In Vitro Diagnostic Medical Device |
|
8. |
Specific evaluation report including Sensitivity & Specificity Test at NABL accredited Lab or central medical device testing laboratory or medical device testing laboratory or medical device testing laboratory registered under sub- rule (3) or rule 83 of MDR 2017 |
|
|
9. |
Specimen batch test report for at least consecutive 3 batches showing specifications of each testing parameter |
|
|
10. |
Stability Test Report-Claimed Stability (Accelerated & Real Time,Shipping Stability and In-Use stability) |
|
|
11. |
Test License or existing regulatory License |
|
|
12 |
Undertaking Letter stating that Compliance with QMS |
|
|
13. |
Labelling and IFU |
|
|
14. |
Fee Challan |
|
|
15. |
Legal Form |
|
The list of documents to be needed to apply manufacturing licence for Class C & D Medical Device and IVD Medical Device-
|
Sl no. |
Medical Device |
IVD device |
|
1. |
Cover Letter |
|
|
2. |
Constitution details of the firm(of domestic manufacturer or authorised agent) |
|
|
3. |
The Establishment/Site ownership/Tenancy Agreement |
|
|
4. |
ISO 13485 Certificate or similar QMS Certificate |
|
|
5. |
Site or Plant master file |
|
|
6. |
QMS Document Quality Manual Quality Policy Control of Documents Control of Records Management Responsibility -Management Review Meeting Internal Audit Procedure for Training Environmental requirement compliance as per Annexure A in Fifth Schedule |
|
|
7. |
Device Master File |
|
|
8. |
Essential Principle Checklist of the Medical Device |
Essential Principle Checklist of the In Vitro Diagnostic Medical Devjce |
|
9. |
Risk Analysis and Control Report |
|
|
10. |
Performance Evaluation Report(Accuracy, repeatability, reproducibility,Sensitivity,Specificity) |
|
|
11. |
Clinical Evaluation Report |
|
|
12. |
Post Market Surveillance Report |
|
|
13. |
Specific evaluation report including sensitivity & Specificity Test at NABL accredited Lab or central medical device testing laboratory or medical device testing laboratory registered under Sub-rule(3) of rule 83 of MDR 2017 |
|
|
14. |
Not Applicable |
NOC from Department of Animal Husbandry,Ministry of Agriculture,In case of Veterinary IVD kits(if applicable) |
|
15. |
Not Applicable |
NOC from Bhaba Atomic Research Centre(BARC),Mumbai,in case Radioimmunoassay Kits(if applicable) |
|
16. |
Stability Test Report- Claimed Stability(Accelerated & Real Time,Shipping,Stability and In-Use Stability) |
|
|
17. |
Test License or existing regulatory License |
|
|
18. |
Undertaking Letter stating that Compliance with QMS |
|
|
19. |
Labelling and IFU |
|
|
20. |
Fee Challan |
|
|
21. |
Legal form |
|
The processes involving or supporting in Manufacturing of medical device and IVD devices is discussed below-
- User account creation
- Apply for Test licence form MD-12
- Grant Test License in Form MD-13
- Documents Preparation for product,site Master file as per Fourth Schedule
- Clinical Evaluation at CDSCO approved Laboratories for Notified Products or NABL accredited Labs for other products
- Prepare document for clinical evaluation,ethical committee clearance
- Apply for clinical performance evaluation of
- Medical Device: Form MD-22
- IVD device :Form MD-24
- CLA will get approval from Central testing laboratories
- Grant approval for clinical performance evaluation of
- Medical device: Form MD-23
- IVD device: Form MD-25
- Apply for Manufacturing or import licence for
- Medical device :Form MD-26
- IVD device:Form MD-28
- CLA will get approval from Central Testing Laboratories
- Grant manufacturing or import license for
- Medical device:Form MD-27
- IVD device: Form MD-29
The resources required for manufacturing medical devices include-
- Ideation and conceptualization
- Site or Plant master file
- Device description
- Personal hygiene requirements
- Work environment
- Temperature control
- Bioburden control
- Humidity control
- Area segregation
- Storage of incoming raw materials or lacking materials.Bulk and finished kits
- Water treatment plant requirements
- Premises and layout
- Clean room
- Control environment
- Lighting
- Specific environment for handling process
- Pest control process
- Water disposal system process
- Power backup-DG
- Preventive maintenance process
- Equipment
- Manufacturing
- Testing
- Storage/preservation
- Sterilization
- Filing
- Packing
- Maintenance
- Calibration(NABL Accredited Centre)
- Cleaning and sanitation
- Testing fascilities
- Production
- List of products
- Production Operations
- Area demarcation
- Process validation
- Sterilization process
- Quality assurance
- Storage
- Temperature controlled environment
- Aseptic condition
- Storage of raw materials,bulk or semi finished and finished products of medical device and In Vitro Diagnostic Device at specified
Information to be covered in the Device Master File of medical devices-
- Executive summary
- Product description, specifications,variants and accessories
- Labeling
- Design and manufacturing information
- Essential Principle Checklist
- Risk analysis and control record
- Verification and validation of medical device
Executive summary
- The executive summary is provided by the manufacturer and it contains
- Product name
- Intended Use
- Manufacturers Information
- Risk classification
- Information on Product(Sterile or non Sterile)
- Device classification as per MDR 2017
- Regulatory status and Product Registration in Other Countries (If applicable)
- class of device, novel features of the device (if any), shelf life of the device and a synopsis on the content of the dossier.
- Similar Products in India
- Risk Analysis and Control Record Document Details
- Declaration of Conformity (DOC) or Design Certificate (if applicable)
- Design Examination Certificate, Declaration of Conformity, Mark of Conformity Certificate, Design Certificate (if applicable). Copy of such certificate(s) shall be enclosed.
- Marketing History Price Details
- List of the regulatory approvals or marketing clearance obtained (submit respective copies of approval Certificates):
|
Country |
Approved Indication |
Approved Shelf life |
Class of Device |
Date of First Approval |
|
USA |
||||
|
Australia |
||||
|
Japan |
||||
|
Canada |
||||
|
European Union |
||||
|
Others* |
*Optional- Status of market clearance pending, rejected or withdrawn
|
Regulatory Agency of the country |
Indication for use |
Registration status and date |
Reason for rejection/ withdrawal, if any |
- Safety & Performance Related information (Adverse Events & FSCA)
- (a) Summary of reportable event and field safety corrective action from the date of introduction:-
- For Serious Adverse Event :
|
S.N. |
Serious Adverse Event (SAE) |
Duration |
Number of the SAE reported |
Total Units sold |
Lot/ Batch No. |
|
|
From |
To |
|||||
- For Field Safety Corrective Action (FSCA):
|
Date of FSCA |
Reason for FSCA |
Countries where FSCA was conducted |
Description of the action taken |
- Device Description or Product Specifications - Available in IFU or Brochures
- Appearance Limitation
- Functional Specifications
- Predicate Device information
- Similar Device available in India
- The device's dossier must include the following descriptive data:
- (a) A general description that includes the product's name in general, the model name, the model number, the materials used in construction, the intended use, the indications, [instructions for use or electronic instructions for use], the contraindications, the warnings, the precautions, and any potential negative effects;
- (b) the intended patient group, the illness that will be identified or treated, as well as additional factors such patient selection criteria;
- (c) the mode of action or operating principle, with animation or movies (if available);
- d) a justification for any unique features;
- (e) a description of any additional medical devices, non-medical products, and accessories that are intended to be used in conjunction with it. It should be made clear whether these additional products are medical devices or accessories.
- (f) a description or comprehensive list of all the different configurations or variants of the device that will be made available;
- (g) a general description of the key functional elements, such as its parts or components (including software if appropriate), its formulation, its composition, and its functionality, and where appropriate, this will include labelled pictorial representations (such as diagrams, photographs, and drawings), clearly indicating key parts or components, including suffi sufficient explanation to understand the drawings and diagrams;
- (h) a description of the materials used in important functioning components and those in direct or indirect contact with a human body, such as when body fluids are circulated outside the body. a thorough chemical, biological, and physical evaluation of the medical device's material(s);
- Information about the radiation source (such as radioisotopes) and the material used to shield from unintentional, stray, or dispersed radiation from patients, users, and other people must be provided for medical devices designed to release ionising radiation.
- 2 Product Specification. The dossier should include a summary of the features, size, and performance characteristics of the medical equipment, as well as any variations and accessories, that are normally available to end users in brochures, catalogues, and other publications.
- 3 Reference to the premise or earlier iterations of the gadget. The dossier should include a summary of: Where applicable to proving adherence to the fundamental principles and to providing broad background information,
- (a) The manufacturer's prior genealogical relationship to the alleged predicate device(s).
- (b) predicate devices available in local and international market; and
- (c) a comparative study to demonstrate the predicate device(sclaimed )'s substantial equivalent.
- Labelling
- Label should contain Products Name,Cat. no/Reference Non,Kit content, individual component label, manufacturers Information
- Refer ISO 15223 for using symbols for Medical Device and IVD
4.Design and Manufacturing information
Manufacturing information
5.Essential Principle Checklist
(i) The dossier should contain the following:
- (a) the essential principles;
- (b) whether each essential principle applies to the device and if not, why not;
- (c) the method used to demonstrate conformity with each essential principle that applies;
- (d) a reference for the method employed (e.g., standard); and
- (e) the precise identity of the controlled document that offers evidence of conformity with each method Used.
(ii) Methods used to demonstrate conformity may include one or more of the following:
- (a) conformity with standards as referred to in rule 7;
- (b) conformity with an in-house test method;
- (c) the evaluation of pre-clinical and clinical evidence:
- (d) comparison to a similar device already available on the market.
A template for a checklist is shown in as under:
|
Essential Principle |
Relevant Yes/ No |
Specification/ standard Sub-clause/ reference |
Complies Yes/ No |
Document Reference Justification and/ or comments |
6.Risk analysis and control summary
Risk:
- Combination of the probability of occurrence of harm and the severity of that harm
Risk management:
- Systematic application of management policies, procedures and practices to the tasks of analysing, evaluating, controlling and monitoring risk
- The Organization shall apply a risk based approach to the control of the appropriate processes
- This risk analysis should be based on prescribed standards and be part of the manufacturer's risk management plan based on complexity and risk class of the device
- The technique used to analyse the risk must be specified, to ensure that it is appropriate for the medical device and risk involved. The risks and benefits associated with the use of the medical device should be described
- The risk analysis submitted shall have periodic updation of the risks identified as per risk management plan
7.Verification and validation studies have undertaken conformity of the device, wherever applicable
(a) Engineering tests;
(b) Laboratory tests;
(c) Simulated use testing;
(d) Any animal tests for demonstrating feasibility or proof of concept of the finished device;
(e) Any published literature regarding the device or substantially similar devices.
Summary information may include-
(i) declaration or certificate of conformity to a recognized standard and summary of the data if no acceptance criteria are specified in the standard;
(ii) declaration or certificate of conformity to a published standard that has not been recognized, supported by a rationale for its use, and summary o the data if no acceptance criteria are specified in the standard;
(iii) declaration or certificate of conformity to a professional guideline, industry method, or in-house test method, supported by a rationale for its use, a description of the method used, and a summary of the data in detail to allow assessment of its adequacy;
(iv) a review of published literature regarding the device or substantially similar devices
Biocompatibility Test
- (i) To characterize the physical, chemical, toxicological and biological response of a material, detailed information should be included on the tests conducted, standards applied, test protocols, the analysis of data and the summary of results. At a minimum, "tests should be conducted on samples from the finished, sterilized (when supplied sterile) device
- (ii) Safety should be assessed in healthy or diseased animal model (intended to be treated by such medical device), as appropriate, demonstrating reaction to active and basic parts of the devices on absolute tissue, local tissue as well as whole organ, clearly recording local, general and systemic adverse reactions, risks or potential risks and performance of device in line with intended use. Wherever possible, histopathology, pathophysiology and path anatomy should be carried out
- (iii) ISO-10993, Biological Evaluation of Medical Devices, should be followed for conducting bio-compatibility study for invasive medical devices should be carried out.
Medicinal Substances
- This dossier should provide detailed information concerning that medicinal substance,its identity and source,the intended reason for its presence, and its safety and performance in the intended application
Biological safety
- Process Validation to demonstrate the safety of Biological substances
- The dossier should contain a list of all materials of animal or human origin used in the device
- Detailed information should be provided concerning the selection of sources or donors; the harvesting, processing, preservation, testing and handling of tissues, cells and substances of such origin
Sterilization
- Provide Sterility Test report to demonstrate the validation of sterilization process
Software Verification and Validation
- Provide information of Software details
- Verification and Validation software interlinked with medical device
Animal Studies
- Animal model have been undertaken to provide evidence of conformity
Stability data
- Provide Real Time and Accelerated Stability study report to demonstrate the claimed Shelf life of the medical device
- Stability study depend on the nature of Device,recommended storage condition and sterility nature
Clinical evidence
- Perform clinical investigation of the device to demonstrate the biocompatibility ,safety and other performance characteristics
Post Marketing Surveillance Data(Vigilance reporting)
- The dossier should contain the Post Marketing Surveillance or Vigilance Reporting procedures and data collected by the manufacturer encompassing the details of the complaints received and corrective and Preventive actions taken for the same
Device disposal information
- Include device disposal information based on the hazardous severity of the Raw Materials as per CLP regulation (EC) No. 1907/2006(EC) No. 1272/2008 and (EU) No. 453/2010
Device Master File for In Vitro Diagnostic Medical Devices
Contents
- Executive Summary
- Product Description, Specifications, Variants, and Accessories
- Essential Principle Checklist
- Risk Analysis and Control Record
- Design and Manufacturing Information
- Verification and Validation of Device
- Analytical Studies
- Specimen Type
- Analytical performance characteristics
- Analytical sensitivity
- Analytical Specificity
- Metrological Traceability
- Measurement Range of Assay
- Definition of Assay Cut-off
- Stability
- Claimed Shelf Life
- In Use Stability
- Shipping Stability
- Clinical Evidence
- Labelling
- Post marketing surveillance data
- Information required to be submitted for the in vitro diagnostic medical device
- Executive Summary
- Product Name
- Intended Use
- Manufacturers Information
- Device Classification as per MDR-2017
- Regulatory status and Product Registration in Other Countries (If applicable)
- Similar Products in India
- Marketing History
- Price Details
- Safety & Performance Related information (Adverse Events & FSCA) Information related to the Device contains Animal or human fluids or derivatives, Cells, tissues, and/or derivatives of microbial recombinant origin
- List of regulatory approvals or marketing clearance obtained in below format (submit respective copy of approval certificate).
|
S.N. |
Name of the country |
Approved indication |
Approved shelf life |
Composition |
Risk Class |
Date of first approval |
- Status of pending request for market clearance
|
Regulatory Agency of the country |
Intended use |
Indication for use |
Registration status and date |
Reason for rejection/ withdrawal, if any |
- Safety and performance related information on the in vitro diagnostic medical device:
- (a) Summary of reportable events and field safety corrective action from the date of introduction.
- For adverse event (false diagnosis or any other hazard during its use)
|
Adverse event (false diagnosis) |
Frequency of occurrence during the period (number of report/ total units sold) |
- For Field Safety Corrective Action (FSCA)
|
Date of FSCA |
Reason for FSCA |
Countries where FSCA was conducted |
Description of the action taken |
- Device Description
- (1) what is detected;
- (2) its function (for example screening, monitoring, diagnostic or aid to
- diagnosis, staging or aid to staging of disease);
- (3) the specific disorder, condition or risk factor of interest that it is
- intended to detect, define or differentiate;
- (4) whether it is automated or not;
- (5) whether it is qualitative or quantitative;
- (6) the type of specimen required (eg. serum, plashna, whole blood,tissue biopsy, urine);
- (7) testing population;
- Product Name, Pack size, Cat No.
- Materials of Construction (Reagent, Assay Controls & Calibrators, Antigens, Antibodies and Nucleic Acids)
- Intended User
- Principle of Operation of the device
- Risk Based Class of the device
- Specimen Collection
- Description of Software and other functional elementS
- Accessories information
- New IVD Device:
- ■ Demonstrate conformity to the essential principles, and to provide general background information, the device master file may provide a summary of Clinical Performance Evaluation reports
- Predicate IVD Device:
- ■Summary of Adverse Event Report, External Certificate or documents that is evident for conformity with essential Principle and Comparative Analysis to prove similarity
- Essential Principle Checklist
(i) The dossier should contain the following:
- (a) The essential principles;
- (b) Whether each essential principle applies to the device and if not, why not;
- (c) The method used to demonstrate conformity with each essential principle that applies;
- (d) A reference for the method employed (e.g., standard); and
- (e) The precise identity of the controlled document that offers evidence of conformity with each method used.
(ii) Methods used to demonstrate conformity may include one or more of the following:
- (a) Conformity with standards as referred to in rule 7;
- (b) Conformity with an in-house test method;
- (c) The evaluation of pre-clinical and clinical evidence;
- (d) Comparison to a similar device already available on the market.
- Risk Management
- The Organization shall apply a risk based approach to the control of the appropriate processes This risk analysis should be based on prescribed standards and be part of the manufacturer's risk management plan based on complexity and risk class of the device.
- The technique used to analyse the risk must be specified, to ensure that it is appropriate for the medical device and risk involved. The risks and benefits associated with the use of the medical device should be described.
- The risk analysis submitted shall have periodic updation of the risks identified as per risk management plan.
5.Design and Manufacturing Information-
- Design Information
- Provide Manufacturer information
- Name and Address
- Competency Available Resources
Manufacturing information
6.Verification and validation studies undertaken to demonstrate conformity of the device
- This Report should contain
- (a) The study protocol;
- (b) The study results;
- (c ) The study conclusion
- Verification phase steps
- Requirement gathering
- System analysis
- Software design
- Module design
- Validation phase steps
- Acceptance testing
- System testing
- Integration testing
- Unit testing
- Execution of Coding
- Analytical Studies
- Analytical Studies should include Sensitivity and specificity of the device Applicable to Quantitative, Qualitative and Semi-Quantitative
- Difference between Analyser and reagents
- Specimen Compatibility Test
This study to demonstrate the device compatible with types of devices, applicable to IVD reagents and Kits
- Serum
- Plasma
- Urine
- CSF
- Other Body Fluids Tissue
- It includes stability of the analytes, specimen storage, transportation, and freeze/thaw cycle
9.Analytical Performance characteristics
Accuracy
- It provides the information on the trueness of the measurement procedure
- Trueness measures apply to both quantitative and qualitative assays only when a certified reference material or certified reference method is available
Reproducibility/Intermediate Precision
- Reproducibility estimates and information about the studies used to estimate,as appropriate,variability between days,runs,sites,lots,operators and instruments
- Such variability is also known as "Intermediate Precision".
10.Analytical Sensitivity
It include information about specimen type and preparation including matrix, analyte (measurand) levels.
For example:
- (a) Number of standard deviations above the mean value of the sample without analyte (measurand), commonly referred to as Limit of Blank (LoB).
- (b) Lowest concentration distinguishable from zero, based on measurements of samples containing analyte (measurand), commonly referred to as limit of detection (LOD).
- (c) Lowest concentration at which precision and/or trueness are within specified criteria, commonly referred to as limit of quantitation (LoQ).
- * For Class C and D in vitro diagnostic medical devices, detailed informationshould be provided.
- Diagnostic sensitivity' means the ability of a device to identify the presence of a target marker associated with a particular disease or condition.
- True Positive Diagnostic Sensitivity (%): ----------------- X 100
- True Positive + True Negative
11. Analytical Sensitivity
It include information about specimen type and preparation including matrix, analyte (measurand) levels.
For example:
- (a) Number of standard deviations above the mean value of the sample without analyte (measurand), commonly referred to as Limit of Blank (LoB).
- (b) Lowest concentration distinguishable from zero, based on measurements of samples containing analyte (measurand), commonly referred to as limit of detection (LOD).
- (c) Lowest concentration at which precision and/or trueness are within specified criteria, commonly referred to as limit of quantitation (LoQ).
- * For Class C and D in vitro diagnostic medical devices, detailed informationshould be provided.
- Diagnostic sensitivity' means the ability of a device to identify the presence of a target marker associated with a particular disease or condition.
- True Positive Diagnostic Sensitivity (%): ----------------- X 100
- True Positive + True Negative
- Metrological traceability of Calibrator and Control Material values Test
It demonstrates the information about methods and acceptance criteria for the metrological traceability to reference materials and/or reference measurement procedures and a description of value assignment and validation.
- Measuring range of the assay Test
- This section should demonstrate the minimum limit of detection and maximum limit of detection (Linearity) of the products Also Describe the hooks effect or prozone effect
- Assay Cut-Off Test
This section should provide a summary of analytical data with a description of the study design including methods for determining the assay cut-off, including:
- (a) the population(s) studied (demographics / selection / inclusion and exclusion criteria / number of individuals included);
- (b) method or mode of characterization of specimens; and
- (c) Statistical methods e.g. Receiver Operator Characteristic (ROC) to generate results and if applicable, define gray-zone/equivocal zone.
- Stability (Excluding specimen stability)
This section should describe claimed shelf life, in use stability and shipping studies.
- Claimed Shelf Life Test
- This section should demonstrate the evident for claimed shelf life when stored at recommended storage condition.
- Accelerated Stability Study will demonstrate when the device exposed at high temperature or humidity depend on the device as per Q10 or Arrhenius equation.
- At least three different lots manufactured under conditions that are essentially equivalent to routine production conditions.
- In Use Stability
- This section should provide information on in use stability studies for one lot reflecting actual routine use of the device (real or simulated).
- This may include open vial stability and/or, for automated instruments, on board stability.
- In the case of automated instrumentation if calibration stability is claimed, supporting data should be included.
- Shipping Stability
- This section should provide information on shipping stability studies for one lot to evaluate the tolerance of products to the anticipated shippingconditions such as extreme heat or cold.
- Clinical Evidence
- The device master file should contain the Clinical Evidence, Evaluation report that demonstrates conformity of the in vitro diagnostic medical device to the Essential Principles that apply to it.
- Labelling
- Label should contain Products Name, Cat.No / Reference No., Kit ,Content, Individual Component Label, Manufacturers information,Symbols for storage, handling, warning.
- Refer ISO 15223 for using symbols for Medical Device and IVD
- Post Marketing Surveillance Study
The dossier should contain the Post Marketing Surveillance or Vigilance Reporting procedures and data collected by the manufacturer encompassing the details of the complaints received and corrective and Preventive actions taken for the same.
- Details of Critical Raw Materials like Antigens, Antibodies and Critical Processes.
- Test Protocol for testing the Device including sensitivity and specificity
- In case of imported diagnostic in vitro diagnostic medical devices, the report of evaluation in details conducted by the National Control Authority of country of origin
- Specimen batch test report for at least consecutive 3 batches
- The detailed test report of all the components used/packed in the finished in vitro diagnostic medical device
- Pack Size and Labeling
- Product inserts / IFU
- Specific evaluation report, if done by any laboratory in India, showing the sensitivity and specificity of the in vitro diagnostic medical device
- Specific processing like safe handling, material control, area control, process control, and stability studies, storage at quarantine stage and finished stage, packaging should be highlighted in the product dossier
- All reports submitted as a part of the dossier should be signed and dated by the responsible person.
- Batch Release Certificates and Certificate of Analysis of finished product for minimum 3 consecutive batches should be oduct for minimu consecutive bat submitted.
- All certificates submitted must be within the validity period. 4. Any information which is not relevant for the subject device may be be stated as 'Not Applicable' in the relevant Sections/Columns of the above format, and reasons for non-applicability should be provided.
22. Information required for New Medical Device and In Vitro Diagnostic Device-
The documents to be needed to apply manufacturing licence for new Medical Device is given below-
- Design Analysis data including, (whichever applicable)
(a) design input and design output documents;
(b) mechanical and electrical tests;
(c) reliability tests;
(d) validation of software relating to the function of the device,
(e) any performance tests;
(f) in vitro tests.
- Bio-compatibility tests data, Report of bio-compatibility tests along with rationale for selecting these tests. Summary report of the biocompatibility study including the conclusion of the study.
- Risk Management data
- Animal Performance study data
- Pilot or Pivotal Clinical Investigation data, including that carried out in other countries if any
- In case, if waiver from clinical investigation is claimed in accordance with the provisions of Medical Device Rules, 2017, the information or device Rules, 2017, the supporting data shall be submitted
- Regulatory status and Restriction on use in other countries (if any) where marketed or approved
- Proposed Instruction for use and labels
The documents to be needed to apply manufacturing license for a new IVD Medical Device is given below-
- Device data including, (whichever is applicable)
(i) design input, design output documents, stability data;
(ii) device specification including specificity, sensitivity, reproducing,reputability;
(iii) product validation and software validation relating to the fur the device (if any);
(iv) performance evaluation report from a laboratory designate sub-rule (1) of rule 19.
- Risk Management data.
- Clinical Performance Evaluation data carried out in India and in other countries (if any).
- Regulatory status and Restriction on use in other countries (if any) where marketed or approved.
- Proposed Instruction for use and labels.
Reference-
http://www.bareactslive.com/ACA/act2713.htm
https://www.emergobyul.com/resources/india-process-chart
https://cdsco.gov.in/opencms/opencms/en/Medical-Device-Diagnostics/
http://www-naweb.iaea.org/napc/iachem/training-modules/Web_PDF_files/SMF_Example_Neptune.pdf