Project 2 - Clinical Investigation Plan
Clinical investigation plan (CIP) means the document that states the rationale, objectives, design and proposed analysis, methodology, monitoring, conduct and record-keeping of the Clinical Investigation and all amendments thereto as the Parties may from time to time agree in accordance with concurrence to the country specific regulation and which have also been signed by the Investigator/ industry involved in it. Such amendments will be signed by the parties involved and form a part of this clinical study trial agreement. The CIP shall be designed in such a way as to optimize the scientific validity and reproducibility of the results of the study in accordance with current clinical knowledge and practice so as to fulfil the objectives of the investigation.
A Clinical Investigation Plan (CIP) is any systematic investigation of a medical device that involves one or more human subjects. The CIP describes the rationale, objectives, design methodology, monitoring, conduct, record-keeping and the method of analysis for a clinical investigation.The CIP shall be a document developed by the sponsor and the clinical investigator(s). The CIP shall be designed in such a way as tooptimise the scientific validity and reproducibility of the results of the study in accordance with current clinical knowledge and practice so as to fulfil the objectives of the investigation.
Key elements of the CIP include:
- •General information to include a comprehensive list of all the CIs, PIs, coordinating CIs, and investigations centers/sites, name and address of the sponsor, monitoring arrangements, data and quality management, an overall synopsis of the clinical investigation, approval and agreement to the CIP
- •Identification and description of the medical device to be investigated
- •Preliminary investigations and justification of the study including literature review, preclinical testing, previous clinical experience, and device risk analysis and assessment (this process is described in EN ISO 14971:2007 which is explored in Chapter 6 “Risk Management”)
- •Objectives of the clinical investigation
- •Decision of the clinical investigation
- •Statistical considerations
- •Deviations from the CIP and how they are handled and recorded
- •Amendments to the CIP (ISO, 2003, p. 6–11)
- •Adverse events (AEs) and adverse device effects
- •Early termination or suspension of the investigation
- •Publication policy
- •Case report forms (CRF) (the means by which data are captured during the investigation)
Types Of Clinical Investigation As Per The Medical Device Regulations 2022
Pilot Clinical Investigation –
A pilot clinical investigation refers to a clinical investigation that is performed on human participants for the first time. Pilot clinical investigations are performed in a small group of patients with the condition or disease being studied to obtain specific information about the device prior to initiating the pivotal clinical study. A pilot clinical investigation is an exploratory study that provides information about the safety and performance of a device but does not provide support for any specific therapeutic or mechanistic claims. A pilot clinical investigation usually has the following objectives:
-
Evaluation of feasibility (e.g., preliminary device performance).
-
Examining eligibility criteria and their application to pivotal controlled investigations, ascertaining possible harm (preliminary Safety Evaluations).
-
Studying device mechanism
-
Validating a method to determine an outcome measure
-
To validate a surrogate outcome, use a defined device method.
-
Evaluation of the logistics of pivotal investigations for performance
Pivotal Clinical Investigation –
Pivotal clinical investigations are definitive or confirmatory study that gathers evidence to support the safety and effectiveness of the medical device intended for use. Pivotal clinical investigations are conducted in a larger population with the condition or disease being studied. Data from the pilot clinical investigation must also be submitted for a pivotal study. A clinical investigation must be conducted in India for any investigational medical device that is being developed in India. Data generated, and a Pilot clinical investigation should be done.
Pivotal studies are required for any investigational medical devices that do not have a predicate device but have been approved for sale or distribution in another country than India. These pivotal studies should be done primarily to show safety and effectiveness in Indian patients when the device is used as directed in the prescribing instructions. To ensure that data from the pilot study is consistent with data generated in India, pivotal clinical investigations on Indian subjects must be conducted before any data from the pilot study can be submitted. The sponsor should submit an annual status report on each clinical investigation to CDSCO, detailing whether it is currently ongoing, completed, or terminated.
Application for Grant of permission to conduct clinical investigation of an investigational medical device IVD
- Biocompatibility study- By definition, a measurement of how compatible a device or material is with a biological system. “Ability of a medical device or a material to perform with an appropriate host response in a specific application”.
- Information specified in table 3 shall be submitted along with investigator's brochure as prescribed in table 4, clinical investigational plan as prescribed in table 5, case report form as prescribed in table 6, serious adverse event reported, if any, as prescribed in table 7, informed consent form as prescribed in table 8, investigator's undertaking as prescribed in table 9, of this schedule and ethics committee approval, if available, as prescribed in appendix VIII of schedule Y of the drugs and cosmetics rules, 945.
- Regulatory status in other countries, if any, on use of investigational medical device in other countries,Prescription based device
Exclusion of certain age groups Warning about adverse device effect Information on withdrawn in any country by the manufacturer or by regulatory authority, such information shall also be furnished along with reasons Sponsor should submit the above information to the Central Licensing Authority during the entire duration of marketing of the said medical device in the Country. - Proposed Instruction for use or direction for use and labels to be submitted along with application. Provided that after submission and approval by the central licensing authority, no change in the instructions for use shall be effected without such changes having been approved by the central licensing authority
- Report of clinical investigation should be in consonance with the format
There are several key policies and guidelines that are typically involved in the development and conduct of a clinical investigation plan. These include:
Good Clinical Practice (GCP) guidelines: These guidelines provide a set of ethical and scientific principles for the design, conduct, monitoring, and reporting of clinical trials. They are designed to ensure that the rights, safety, and well-being of study participants are protected, and that the data generated from the study are reliable and of high quality.
International Conference on Harmonisation (ICH) guidelines: These guidelines provide a set of standards for the design, conduct, and reporting of clinical trials that are conducted across different countries. They are designed to facilitate the global development and registration of new medical products, and to ensure that the results of clinical trials are consistent and comparable.
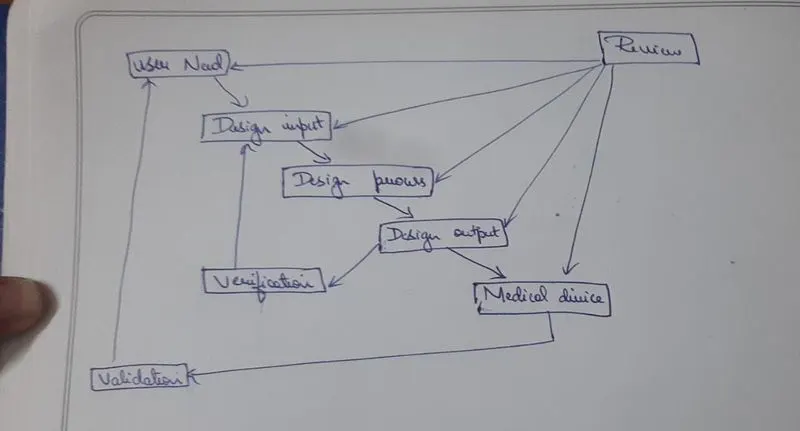
Management, creation and execution of a design history file for each type of device take experience and grit. Completing a risk analysis, or hazard analysis early in the design phase for medical devices is not only required, but is important for successful development. The FDA led in this respect, creating regulations documented in 21 CFR Section 820. Today we have standards such as ISO 13485 and ISO 14971 (risk analysis) to guide our global de-risking of medical devices. We have developed industry standard best practices for design control management including risk analysis processes that help move a device efficiently and effectively through the early stages of development. The regulations and standards are straightforward to implement, and gain complexity as company size and risk tolerance grow. Keeping control or iterating on your quality management system of industry standard best practices is important when developing a medical device and requires surveillance and dedication to keeping patients safe while not being sidetracked. Today, as our industry has evolved, device risk requirement generation has become a part of medical device design and development. These tools are useful and necessary in the development process. We have experience with risk analysis, creation of user needs, design control file creation, process validation, clinical evaluations and many other activities as identified in ISO 14971 that are required to produce a safe medical device. We can help at any stage with the development of your medical device.
Institutional Review Board (IRB) guidelines: An IRB is an independent committee that reviews and approves clinical research studies involving human subjects. IRBs are responsible for ensuring that clinical research studies are conducted in accordance with ethical principles and guidelines, and that the rights and welfare of study participants are protected.
In terms of the justification for a clinical investigation, it is important to clearly articulate the rationale for the study, including the specific research question or hypothesis being tested, and how the study will contribute to the scientific knowledge base. This may include an overview of relevant literature, an explanation of the current state of knowledge in the field, and a description of the potential benefits and risks of the study.
In terms of conducting the investigation, it is important to follow the protocols and procedures outlined in the clinical investigation plan. This may include recruiting and enrolling participants, administering interventions or tests, collecting and managing data, and analyzing and interpreting the results. It is also important to carefully document all aspects of the study, including any deviations from the protocol, and to report the results in a transparent and accurate manner.
It is important to carefully consider the risks and benefits of an investigational medical device and clinical investigation when designing and conducting a study. Risks may include any potential harm or discomfort that may be experienced by study participants, as well as any potential risks to the scientific integrity or validity of the study. Benefits may include the potential for improving patient outcomes or advancing scientific knowledge, as well as any potential economic benefits.
- To visualize and frame the risks and benefits of the investigational medical device and clinical investigation, it may be helpful to create a risk-benefit matrix. This is a tool that allows you to identify and assess the risks and benefits of the study, and to determine whether the potential benefits justify the potential risks.
- To create a risk-benefit matrix, you will need to identify the specific objectives and hypotheses of the study, as well as the potential risks and benefits that may be associated with the investigational medical device and clinical investigation. You can then use this information to evaluate the overall balance of risks and benefits, and to determine whether the study should proceed.
- In some cases, it may be necessary to modify the study design or protocol in order to mitigate or eliminate certain risks, or to maximize the potential benefits of the study. For example, you may need to implement additional safeguards to protect the rights and welfare of study participants, or to improve the scientific validity of the study.
- Ultimately, the decision to proceed with a clinical investigation should be based on a careful assessment of the risks and benefits, and on a determination that the potential benefits justify the potential risks. It is important to carefully consider all of the relevant factors, and to consult with relevant stakeholders, such as study participants, regulatory agencies, and ethics committees, before making a final decision.
- Statistical considerations are an important aspect of the design of a clinical investigation, as they help to ensure that the study is adequately powered to detect any differences or associations that may exist between the experimental and control groups.
There are several key statistical considerations that should be taken into account when designing a clinical investigation:
- Sample size: The sample size is the number of participants in the study. It is important to determine the appropriate sample size based on the statistical power of the study, which is the probability that the study will detect a true difference between the experimental and control groups. A larger sample size increases the statistical power of the study, but also increases the cost and resources required to conduct the study.
- Statistical analysis plan: A statistical analysis plan is a detailed description of the statistical methods that will be used to analyze the data from the study. It should specify the primary and secondary outcomes that will be measured, the statistical tests that will be used, and any assumptions or limitations of the analysis.
- Statistical significance: Statistical significance is the probability that a difference or association between the experimental and control groups is due to a real effect, rather than chance. A commonly used threshold for statistical significance is p < 0.05, which means that there is a less than 5% chance that the difference is due to chance.
- Confidence intervals: Confidence intervals provide a range of values within which the true population value is likely to fall, based on the sample data. They help to provide a sense of the precision of the estimate, and can be used to determine whether the difference between the experimental and control groups is statistically significant.
- Data management is also an important aspect of clinical investigation, as it involves the collection, storage, and analysis of the data generated from the study. This may include establishing processes for collecting and recording data, developing a database or other data management system, and ensuring that the data are accurate, complete, and secure. It is important to have robust data management practices in place in order to ensure the integrity and reliability of the study results.
Statements of compliance refer to the measures and procedures that are in place to ensure that a clinical investigation is conducted in accordance with relevant regulations and guidelines. These may include measures related to the informed consent process, the reporting of adverse events, and the handling of data and other sensitive information.
Informed consent is a process by which study participants are provided with information about a clinical investigation and are given the opportunity to voluntarily agree to participate in the study. The informed consent process is a critical aspect of the overall clinical research process, as it helps to ensure that study participants are fully informed about the potential risks and benefits of the study, and that they are able to make an informed decision about whether or not to participate.
The informed consent process typically involves providing study participants with written information about the study, including the purpose of the study, the procedures that will be performed, the potential risks and benefits of participation, and any alternatives to participation. It also involves giving study participants the opportunity to ask questions and to discuss their concerns with the research team.
In addition to obtaining informed consent from study participants, it is also important to ensure that the clinical investigation is conducted in compliance with all relevant laws, regulations, and guidelines. This may involve obtaining approval from an Institutional Review Board (IRB) or other regulatory body, and following specific protocols and procedures to ensure the ethical conduct of the study.
In terms of adverse events associated with the device, it is important to carefully monitor and document any adverse events or adverse reactions that occur during the course of the study. Adverse events are any negative or unintended effects that occur as a result of the investigational device or the study procedures, and they may range from mild side effects to serious injuries or deaths. It is important to report all adverse events to the appropriate regulatory authorities, and to take appropriate action to mitigate any risks to study participants.
Inclusion and exclusion criteria are used to specify the types of individuals who are eligible or ineligible to participate in a clinical investigation. These criteria are typically based on factors such as age, gender, medical history, current health status, and other characteristics that may affect an individual's ability to participate in the study or the interpretation of the study results.
Inclusion criteria are the criteria that must be met in order for an individual to be eligible to participate in a clinical investigation. These criteria are used to define the target population for the study and to ensure that the study is focused on a specific group of individuals who are most likely to benefit from the investigational product or intervention being tested.
Reasons for clinical Trial Termination
low accrual rate is the biggest reason for trial termination, it is not surprising that the average enrollment efficiency (the number of subjects planned divided by the number of subjects actually enrolled) in these trials for every phase is below 40%. The lowest enrollment efficiency in any phase occurred in Phase III at 32.1%, and the highest occurred in Phase IV at 39.8%. The average number of patients enrolled also suffered, ranging from 19.1 in Phase I up to only 79.9 in Phase III.
This data underscores the need for addressing issues of poor enrollment in clinical trials, especially in Oncology indications, which were the most often terminated therapy area, with low accrual being the reason for this. Clinical trial feasibility planning to minimise risk, account for competitive activity, and ultimately select the right geographic location, sites, and investigators for a trial are paramount.
Conclusion:
Guidelines for early termination of a clinical trial should be established before any data review is undertaken. Interim data analyses in conjunction with the totality of available evidence provides the necessary framework from which Data and Safety Monitoring Boards can make informed and prudent recommendations. Controlled clinical trials should not be prematurely terminated for trivial reasons. Controlled clinical trials should not be terminated prematurely on economic grounds, particularly if the information to be gained adds substantially to the knowledge base on the therapy of disease states. Such, for example, is the case with the early stoppage of the CONVINCE trial, where the motives for its discontinuation were a presumed fiduciary responsibility to the stockholders since it was believed that its findings might not fall into the “blockbuster” category. Finally, continuation of a trial deemed to be futile is wasteful of resources and potentially unethical. This is the case in the ALLHAT study, wherein there was early termination of the doxazosin treatment limb because of an unacceptably high rate of CHF when doxazosin-treated patients were compared to those receiving chlorthalidone. Similarly, there was an early termination of the amlodipine treatment limb of the AASK trial. When amlodipine treated patients were compared to those receiving the ACE inhibitor ramipril there was a sufficient difference in rate of decline of renal function in proteinuric subjects that the Data and Safety Monitoring Board felt that discontinuation of the amlodipine treatment limb to be warranted.